Key Takeaways
- A virtual data room for pharma M&A holds regulatory submissions, clinical data, patent files and manufacturing records
- Valuation often rests on the patent estate and the clinical package rather than on revenue
- Clinical data carries patient privacy obligations that follow it through diligence
- Folder structure should follow the asset, since biotech deals are frequently about one or two programs
- No data room makes you FDA 21 CFR Part 11 compliant. That obligation stays with the sponsor
- Staged access matters more here, because most bidders will see your science and not proceed
- The same room supports licensing and partnership deals, which outnumber acquisitions in this sector
Table of Contents
ToggleA virtual data room for pharma M&A is a permissioned workspace holding the documents a life sciences transaction turns on: regulatory submissions such as INDs and NDAs, clinical trial data, patent and licensing files, manufacturing records, and preclinical results. It differs from a general deal room in what it holds, who reviews it, and the privacy obligations attached to the data.
The difference starts with what is being bought. Most acquisitions value a business. A biotech acquisition frequently values a molecule, a patent estate, and a clinical dataset that may or may not survive its next readout.
That shifts the whole exercise. The reviewers are patent counsel, regulatory specialists and clinical scientists rather than accountants. The crown jewels are documents rather than facilities. And the material that proves the asset is the same material that would let a competitor replicate it.
Why Pharma and Biotech M&A Needs a Data Room
Transactions involving life science, biotech and pharmaceutical companies are complex, with hundreds and sometimes thousands of confidential documents shared among licensing partners, bidders, investors, CROs and regulators, alongside their legal, finance, accounting and leadership teams.
Diligence here concentrates on intellectual property rather than on operations. In many biotech deals the valuation of the IP is effectively the valuation of the entire company, which makes the timely, controlled presentation of those documents central to whether a transaction closes.
These deals also happen rarely enough that few companies have a repeatable internal process for them. When a transaction becomes imminent, the room is usually built from scratch under time pressure, which is where structure decisions get made badly.
What Goes Into a Life Sciences Data Room
The document set differs enough from a standard deal that a generic folder template will not fit it.
| Category | Contents | Primary reviewer |
|---|---|---|
| Intellectual property | Patent families, prosecution history, freedom-to-operate analyses, licensing agreements, assignments | Patent counsel |
| Regulatory | IND and NDA submissions, FDA correspondence, drug master files, marketing applications filed or approved | Regulatory affairs |
| Clinical | Trial protocols, results, safety data, CRO agreements, investigator agreements | Clinical scientists |
| Preclinical | Toxicology, pharmacology, study reports supporting existing or pending products | Scientific diligence |
| Manufacturing | Process documentation, supply agreements, CMO contracts, quality records | Technical operations |
| Commercial and corporate | Financials, cap table, employment agreements, material contracts | Financial and legal |
Two structural points follow from that table. Reviewers are specialists who need their own material and rarely need anyone else’s, so access should be scoped by discipline. And in a company with several programs, the folder tree should follow the asset first and the document type second, because a buyer acquiring one program should not be reading the file on another.
The live page lists five IP document types and stops. That list is retained above and extended across the full set a life sciences buyer actually requests.
Compliance: Where the Obligation Actually Sits
This is where pharma data room marketing tends to overreach, and where a regulatory affairs reader will judge the page. Four frameworks come up, and none of them works the way a compliance badge implies.
FDA 21 CFR Part 11
Part 11 sets the conditions under which the FDA accepts electronic records and electronic signatures as equivalent to paper. It applies to records a company creates, modifies, maintains or transmits to satisfy an FDA requirement, and the FDA guidance on scope and application narrows that scope considerably from how it is often described.
The obligation belongs to the regulated company. A sponsor validates its systems for intended use, controls access, maintains audit trails and defines its procedures. A vendor supplies capabilities that support that work. No software is Part 11 compliant by itself, and any provider claiming otherwise is describing something the regulation does not contain.
For a diligence room the practical question is narrower still: are the regulatory records you are sharing subject to Part 11 in their system of record, and does moving copies into a data room change your obligations there. That is a question for regulatory affairs, not for a vendor.
HIPAA
Clinical trial data containing identifiable patient information falls under HIPAA when handled by a covered entity or its business associate. The relevant vendor question is whether the provider will sign a business associate agreement, since sharing protected health information without one is a violation regardless of encryption strength.
Most sponsors de-identify clinical data before diligence, which changes the analysis. Whether de-identification meets the standard is a decision your privacy counsel makes, not one the data room makes for you.
GDPR
European trial sites mean European data subjects, and personal data belonging to them travels under GDPR wherever your company sits. You are the controller and the provider is a processor. Their certifications support your diligence obligation without discharging it.
GxP
Good laboratory, clinical and manufacturing practice govern how research and production are conducted, not how a deal is run. A diligence data room sits outside those processes. Where GxP intersects with a transaction is in the records themselves: a buyer reviews GxP documentation to assess whether the target’s quality systems will withstand inspection.
| Common Mistake
Treating a vendor certification as your compliance. The FDA’s own Part 11 guidance frames validation around intended use by the regulated party. A provider can hold every certification available and your obligation is unchanged. |
Protecting IP and R&D During Diligence
Here is the tension specific to this sector. To value your asset, a buyer needs to see the science. Most buyers who see it will not proceed, and some of them are running competing programs in the same indication.
That is a different exposure from a standard deal, where a bidder who walks away holds financial data that ages quickly. Preclinical results and formulation detail do not age. They tell a competitor what worked and, just as usefully, what did not.
Three controls address it, and they work together rather than individually.
- Staged disclosure. Early rounds see summary data and patent numbers. Full trial datasets and formulation detail wait for signed terms and a narrowed field
Rights management that survives download. FileProtect restricts printing and forwarding, and revokes access to files a withdrawn bidder has already taken
- Watermarking that identifies the viewer. It prevents nothing directly. It makes a photographed screen traceable, which changes behavior in a way a technical block does not
Trade secrets deserve separate handling from patents, because the protection depends on the information staying secret. Our guide to trade secret protection during M&A covers that distinction, and IP due diligence covers what a buyer examines.
| Worth Knowing
Breaches involving a third party reached 48% in Verizon’s 2026 Data Breach Investigations Report, up 60% year over year. In a biotech process, every bidder, CRO and adviser you admit is a third party holding your preclinical results. |
Beyond Acquisitions: Licensing, Partnerships and Fundraising
Acquisitions are the minority of life sciences transactions. The same document set supports several deal types, and the access model changes between them.
| Deal type | What the counterparty reviews | Access consideration |
|---|---|---|
| Acquisition | The full package across all programs | Broadest disclosure, staged by round |
| IP licensing | One asset, its patent estate and supporting data | Scoped tightly to the licensed program |
| Strategic partnership | The program plus manufacturing and commercial capability | Often reciprocal, with both sides disclosing |
| Fundraising | Program status, regulatory path, use of proceeds | Lightest, and the widest number of viewers |
Licensing deals in particular reward a well-structured room, because the counterparty is evaluating one asset and should see only that asset. Our guide to IP licensing covers how those agreements are structured.
Where AI Genuinely Helps, and Where It Does Not
AI-assisted review is being marketed heavily into life sciences diligence, and the honest position is narrower than the pitch.
What works: organizing large scanned document sets, extracting key terms from contracts at volume, and flagging candidate passages for redaction across thousands of pages. Those are pattern tasks over structured text, and they save real time on a clinical or regulatory file set.
What does not: judgment on scientific validity, on freedom to operate, or on whether a trial result supports a valuation. Those remain specialist work, and a tool presenting an extraction as an assessment is a risk rather than an accelerant.
One further caution specific to this sector. Submitting confidential clinical or formulation data to a general AI platform is a disclosure. Verizon reports shadow AI as the third most common non-malicious insider data-loss event, up fourfold year over year. A room that prevents copying out of the viewer prevents that particular route.
Why CapLinked for Pharma and Biotech Deals
Mapped to the requirements above. CapLinked maintains a dedicated life sciences data room for biotech and pharmaceutical transactions.
On scoped access, permissions are set per folder with files private by default, so patent counsel, regulatory affairs and clinical reviewers each work their own material within one structure. On the question cycle, EZ Q&A routes each query to the specialist who can answer it and keeps the thread attached to the document, which matters when a single trial protocol draws questions from four disciplines.
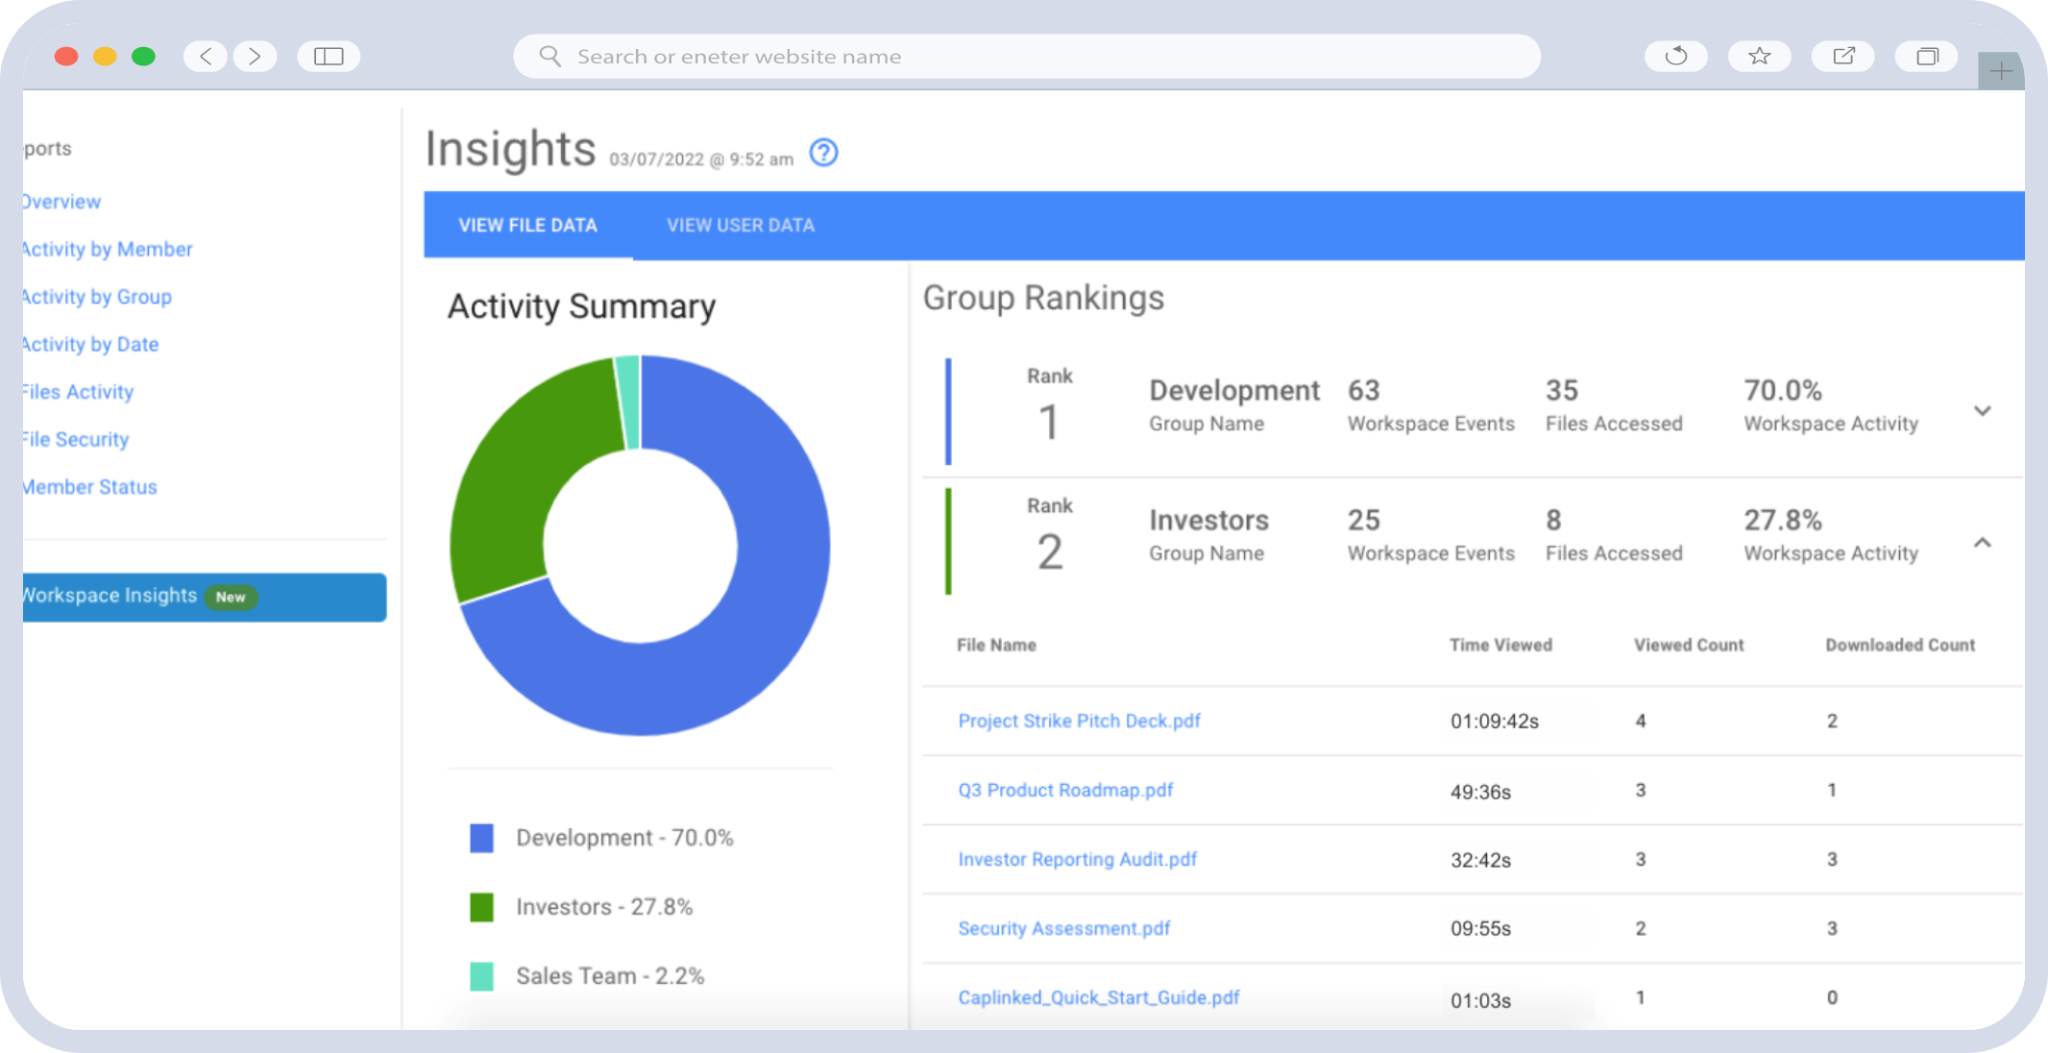
On IP exposure, FileProtect restricts printing and forwarding and revokes access after download, and watermarks carry the viewer’s identity across every page. On the record, the audit trail archives every document interaction in a workspace and exports, which is what supports a sponsor’s own control obligations rather than replacing them.
On the platform itself: 256-bit AES at rest, TLS in transit, independent SOC 2 assessment alongside ISO 27001, PCI SAQ-D and FISMA, and HIPAA and HITECH requirements met including support for signing business associate agreements. The security detail separates which claim is which, and pricing is published rather than quote-gated.
| Running diligence across patent counsel, regulatory affairs and clinical reviewers at once? |
Pharma M&A Data Room FAQ
Why does pharma M&A need a virtual data room?
Life sciences deals turn on IP, clinical data and regulatory submissions reviewed by specialists who each need different material. A permissioned room scopes access by discipline and controls what leaves with bidders who withdraw.
What documents go in a pharma M&A data room?
Patent families and prosecution history, IND and NDA submissions, FDA correspondence, drug master files, clinical protocols and results, preclinical study reports, manufacturing and CMO agreements, plus standard corporate and financial records.
Does a data room make us FDA 21 CFR Part 11 compliant?
No. Part 11 obligations sit with the regulated company, which validates its systems for intended use and defines its procedures. A data room can support those controls through audit trails and access management.
How is clinical trial data protected during diligence?
Through de-identification before sharing where possible, a business associate agreement where protected health information is involved, scoped access limited to clinical reviewers, and rights management that revokes access after a bidder withdraws.
How do you protect IP while letting a buyer evaluate it?
Stage the disclosure. Early rounds see summary data and patent numbers; full datasets and formulation detail wait for signed terms. Combine that with watermarking and rights management that survives download.
Can the same data room support licensing deals?
Yes, and licensing benefits most from good structure. A licensee evaluates one asset and should see only that program, which a folder tree organized by asset handles directly.
Does AI speed up life sciences due diligence?
For organizing large document sets, extracting contract terms and flagging redaction candidates, yes. Scientific validity, freedom to operate and valuation judgment remain specialist work no tool currently replaces.